A toxicology review team at the Food and Drug Administration can open an electronic dataset and view individual animal body weight changes across forty-two days within three keystrokes. When a sponsor submits that exact trial as an eight-hundred-page static text report, the reviewer must manually extract tabular rows and reconcile missing observation points. This procedural friction led regulatory bodies to construct strict electronic data gates.
With more than 20 years of hands-on experience setting up large animal models for regulatory-facing preclinical programs, our team at Biotech Farm has seen firsthand how an electronic technical rejection at the submission gateway halts evaluation before scientific review even begins. For a biotech team with limited runway, an unexpected delay burns capital and pushes planned clinical milestones back.
Clean datasets protect submission timelines. Nonclinical study submissions require standardized electronic structures under published agency frameworks. This guide breaks down dataset expectations, architecture requirements, and operational methods required to deliver compliant nonclinical data packages on schedule.
The single most common — and most preventable — cause of technical rejection we encounter is not bad science. It is improper file naming. A body weight file labeled bw_study101_final.xpt instead of bw.xpt will fail automated validation at the agency intake threshold before any human reviewer ever sees your data. Agencies reject improper filenames at the gateway, regardless of the quality of the underlying science.
What is a SEND dataset in a preclinical submission?
A Standard for Exchange of Nonclinical Data package represents animal study results in a structured tabular format. The system translates laboratory observations into standardized domain files that computer algorithms can query and verify instantly.
Instead of relying on narrative summaries, reviewers load these uniform tables directly into agency screening tools to cross-reference findings across dose groups. The underlying standard was created by the CDISC organization for nonclinical research to mirror the success of human trial data models. Each dataset stores single observations alongside defined animal identifiers, collection intervals, and standardized measurement units.
The standard currently covers single-dose and repeat-dose toxicity as well as carcinogenicity and safety pharmacology studies. Converting raw observations into this model produces clean data records across every evaluation phase.
When are SEND datasets required for regulatory submission?
Electronic datasets become mandatory based on study initiation dates published in the binding agency catalog. The United States Food and Drug Administration specifies these requirements within the official FDA study data submission guidance for human drug development programs.
Studies initiated after December 2016 for commercial applications generally require standardized datasets if the study design falls within an active implementation guide. Commercial Investigational New Drug submissions initiated after December 2017 must also deliver compliant tables for in-scope toxicology work.
Submitting an unstandardized study when rules mandate electronic files leads to immediate rejection at the intake portal. Regulatory staff will not evaluate the study pharmacology until technical criteria are met — meaning your science never even reaches a reviewer’s desk.
What files are mandatory in a SEND dataset preclinical submission package?
A complete submission package contains three distinct file formats working in coordination. Omitting any single file breaks the package structure and invalidates the submission.
1. Domain Data Files (.xpt)
Domain data files formatted in the SAS transport version 5 specification. These files carry the file extension xpt and hold the actual observational records collected during the study.
2. Define.xml Metadata File
A define.xml metadata file that maps every domain variable, controlled terminology codelist, and computational algorithm. This is the master roadmap regulators read first.
3. Nonclinical Study Data Reviewer Guide
An nSDRG provided as an accessible PDF document. This guide explains study conduct deviations, software exceptions, and sponsor decisions that lie outside the standard metadata code.
What is define.xml and why is it critical for submission?
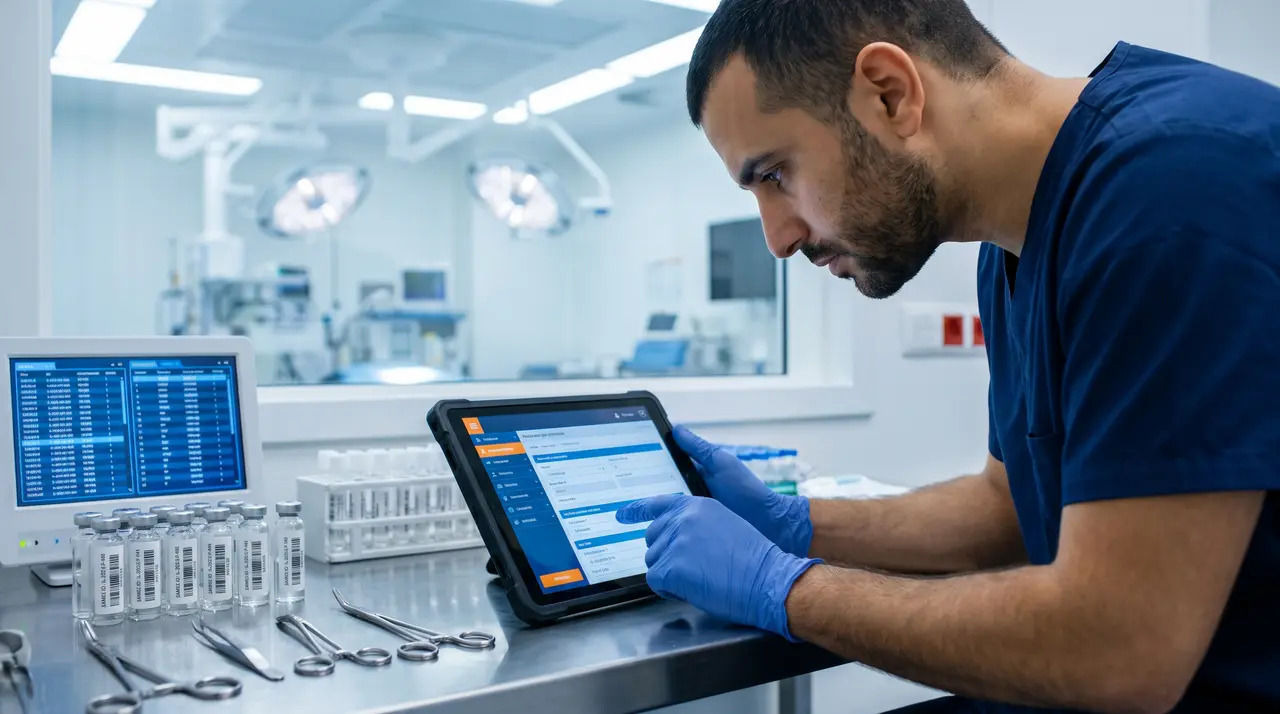
The define.xml file functions as the master roadmap for every table submitted in the package. Regulatory evaluation systems read this file before attempting to parse individual domain records.
This file defines variable structures, indicates expected data types, and references established standard dictionaries. If an analytical tool encounters an unexpected field or an undefined unit, it queries the metadata file for clarification. A missing or corrupt metadata document stops agency review scripts from executing properly. Generating an accurate define document requires complete synchronization between your study documentation and the actual rows inside your data tables.
What is nSDRG and when is it used?
The Nonclinical Study Data Reviewer Guide supplies human context that machine-readable code cannot convey. Standard tables capture numerical readings and categorical findings, but they rarely explain why a specific animal missed an interim blood draw.
This narrative companion describes the conversion process, identifies software tools used, and documents known validation warnings. When a validation script flags an apparent data anomaly, the reviewer consults this document to understand the underlying experimental cause. A well-constructed guide prevents unnecessary information requests from regulatory review divisions during critical evaluation windows.
Where do SEND datasets go within the eCTD structure?
Dataset placement inside the electronic Common Technical Document structure follows strict hierarchy rules. Misplaced files cause gateway indexing scripts to fail during automated intake screening.
Data tables belong in Module 4 under the specific nonclinical study directory. The precise path follows the sequence m4/datasets/study-number/tabulations/send without variation. The corresponding study report sits alongside in the analysis folder within the same section. Regulatory specifications prohibit adding custom folder branches or arbitrary organizational layers inside this designated path.
How should XPT files be named to avoid technical issues?
Transport file naming conventions follow an uncompromising standard designed for automated indexing engines. Every file must match its standardized domain acronym followed strictly by the lowercase xpt extension.
A body weight file must be named bw.xpt without exception. Adding study identifiers, date stamps, or version indicators creates immediate ingestion errors. A file labeled bw_study101_final.xpt will fail automated validation checks at the agency intake threshold.
Regulatory gateways reject improper filenames.
What is the difference between SEND and SDTM?
Both frameworks originate from the Clinical Data Interchange Standards Consortium and share foundational data design principles. However, their internal architectures address entirely different biological models.
The Study Data Tabulation Model supports human clinical trials and tracks informed consent, adverse events, and healthcare encounter records. The nonclinical standard models animal housing environments and maternal-fetal observations as well as terminal necropsies and organ pathology findings. Variables within animal tables capture findings that have no logical equivalent in human clinical hospital systems.
What types of studies are typically in scope for SEND?
The mandate applies primarily to toxicology and safety testing designs that evaluate general systemic tolerance. Single-dose and repeat-dose toxicology as well as lifetime carcinogenicity studies represent the primary study classes subject to enforcement.
Specialized testing providers such as Charles River detail these requirements in a nonclinical data overview for sponsors preparing regulatory filings. Certain safety pharmacology protocols evaluating respiratory or cardiovascular parameters also fall within current scope expectations. Pharmacokinetic studies with systemic bioanalytical sampling intervals require electronic records under current implementation rules.
Is SEND also required for non-GLP studies?
Good Laboratory Practice certification does not determine whether electronic datasets are required. Regulatory applicability depends strictly on the intended use of the trial within the marketing or investigational application.
If an early non-GLP repeat-dose study serves as key safety evidence supporting clinical dose selection, the agency expects standardized electronic tables. Conversely, exploratory discovery trials executed without regulatory submission intent remain entirely exempt from standard tabulation rules. Program sponsors must assess the regulatory objective of each study rather than relying on its laboratory compliance classification.
What is traceability and why is it important in SEND?
Traceability describes the unbroken operational thread connecting raw observation values to the summarized calculations in the final study report. A reviewer must be able to follow every calculation backward to the individual animal record.
If the final study report states an average liver weight of 450 grams for high-dose males, the underlying organ dataset must yield that exact number. Discrepancies between tabulated values and published narrative reports raise regulatory concerns regarding overall data integrity. Ensuring complete traceability requires verified calculation scripts and consistent decimal rounding rules across all project data streams.
A reviewer must be able to follow every calculation backward to the individual animal record. Discrepancies between tabulated values and narrative reports raise concerns about overall data integrity.
— Biotech Farm Study Operations Team
What are common technical rejection criteria for SEND submissions?
Agencies employ automated screening filters called Technical Rejection Criteria to block non-compliant files before human reviewers see them. These checks verify that mandatory components exist and conform to published technical parameters.
Missing a trial summary domain triggers an automatic rejection code during initial screening. Similarly, submitting an eCTD section with a PDF study report but missing the required datasets directory halts the submission. Validation errors in the define.xml schema or mismatched study identifiers across domains also generate catastrophic gateway intake failures.
How do you validate SEND datasets before submission?
Dataset validation requires executing standardized conformance checking software against the complete package. These programs evaluate every column and record against published rules defined by CDISC and regulatory agencies.
Validation engines flag issues ranging from minor terminology discrepancies to major structure violations. Sponsors must inspect each generated warning, correcting actual data errors and documenting legitimate scientific anomalies inside the reviewer guide. Third-party guides such as the Allucent guide to dataset compliance emphasize resolving high-severity rule violations before transmitting packages to regulatory gateways.
What is the CDISC SEND Implementation Guide?
The implementation guide serves as the definitive engineering manual for constructing compliant nonclinical datasets. It defines allowable domain structures, specifies required field names, and establishes structural relationships between related tables.
Different versions of the guide expand coverage to additional nonclinical study types. Version 3.0 focused primarily on standard repeat-dose toxicity, while the 3.1 release added reproductive toxicity and expanded cardiovascular safety domains. Sponsors must ensure their conversion workflows align with the specific version recognized in the active regulatory standards catalog at study initiation.
What role do Controlled Terminologies play in SEND datasets?
Controlled terminologies ensure that scientific terms remain uniform across independent laboratories and study reports. Without standardized dictionaries, one facility might record a tissue observation as enlarged liver while another records hepatic hypertrophy.
The standard relies on terminology maintained by the National Cancer Institute to standardize diagnostic concepts, anatomical sites, and dosage units. Using unauthorized synonyms or non-standard abbreviations triggers immediate validation rule errors. Any necessary custom terms must be formally defined inside the define.xml file as extended terminology.
Operational challenges in real-world SEND conversion
Bioanalytical data conversion friction
Converting twenty-eight days of continuous pharmacokinetic data from sixty laboratory animals exposes several operational friction points. Laboratory information systems frequently output data in wide-table orientations that place timepoints across horizontal column headers.
The standard requires a normalized vertical structure where each individual plasma draw occupies a separate data row with standardized time variables. Transforming these laboratory tables involves mapping sampling intervals, dilution factors, and lower limits of quantification into the pharmacokinetics domain. Inconsistencies between laboratory sample identifiers and in-life animal numbers create validation conflicts that require tedious manual correction.
Large animal study protocols and electronic data capture alignment
Translating surgical observations from large animal models into standard domains introduces distinct data management demands. When conducting medical device safety evaluations in sixty-kilogram domestic swine models, researchers collect complex hemodynamic data alongside standard toxicology parameters.
Facilities like Biotech Farm address this complexity by structuring surgical and physiological recording forms to align with standard domain definitions from study inception. Capturing baseline vitals, anesthesia depths, and post-operative observations using controlled nomenclature eliminates the need for speculative data mapping months later. Structured operational documentation maintains clean records for animal welfare monitoring and regulatory data generation simultaneously.
By structuring recording forms to align with standard domain definitions from study inception, our teams eliminate the speculative data mapping that plagues programs months later. Controlled nomenclature captured in real time means clean records for animal welfare monitoring and regulatory data generation simultaneously.
In-house imaging and specialized telemetry data translation
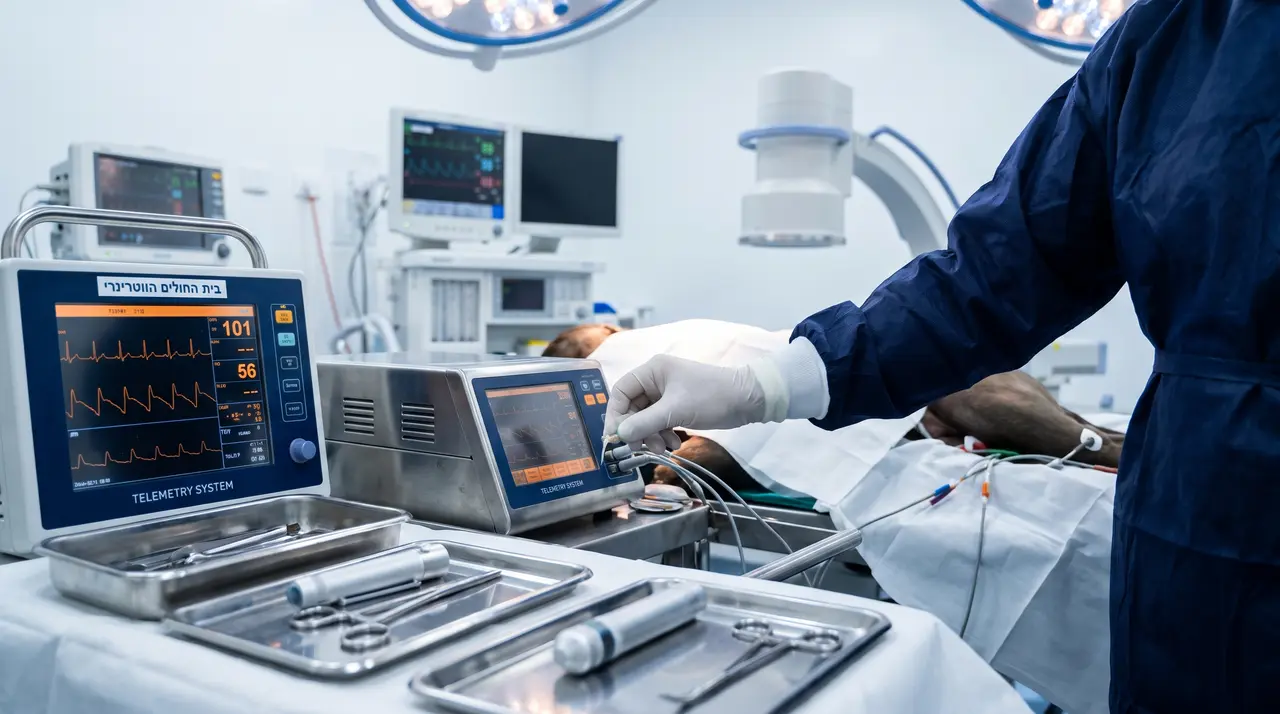
Modern translational research relies on real-time fluoroscopic imaging, high-definition echocardiography, and multi-lead cardiovascular telemetry. Capturing these high-frequency physiological measurements in large animal models yields extensive volumes of raw operational data.
At Biotech Farm, research teams employ specialized surgery suites equipped with fluoroscopy and advanced ultrasound tools to record cardiovascular endpoints cleanly. Structuring these quantitative diagnostic metrics during the active intervention allows technicians to populate cardiovascular domains without data loss. Direct electronic recording avoids the transcription errors common when translating unstructured surgical notes into rigid submission formats.
Protocol design collaboration and raw data governance
Achieving compliance requires continuous alignment between the sponsor, the study director, and the data conversion team throughout the study lifecycle. If protocol amendments modify dosage frequencies or add unscheduled tissue harvests, data collection systems must update corresponding trial summary parameters immediately.
Biotech Farm emphasizes scientific guidance and transparent operational communication to keep study protocols synchronized with final data delivery expectations. Experienced study managers document procedural deviations as they happen rather than reconstructing events during final report publishing. This continuous documentation model simplifies compiling the nonclinical study data reviewer guide and protects project delivery timelines.
Early planning eliminates conversion panic.
Regulatory review stages and data delivery checkpoints
Navigating the nonclinical data lifecycle requires structured verification at every operational milestone. The following breakdown illustrates the primary checkpoints connecting study execution to regulatory submission.
| Project Phase | Operational Objective | Primary File Deliverable | Primary Compliance Risk |
|---|---|---|---|
| Protocol Setup | Align parameters with standardized domains | Trial Design Datasets | Missing required study trial design variables |
| In-Life Testing | Capture raw measurements with standard terms | Raw Instrument Records | Uncontrolled observation terms and missing units |
| Data Mapping | Transform measurements into transport domains | XPT Domain Files | Traceability loss between raw data and tables |
| Package Assembly | Generate metadata and narrative documentation | Define.xml and nSDRG | Validation errors and unexplained study warnings |
| Gateway Filing | Transmit package through submission portal | Complete Module 4 Folder | Technical rejection due to improper folder paths |
Executing systematic checks across each milestone prevents the accumulation of technical debt that can derail formal filings. Regulatory reviewers expect each phase to demonstrate complete operational continuity.
Case Study: Medical Device Safety Evaluation in Swine Models
The challenge: A sponsor needed a medical device safety evaluation in sixty-kilogram domestic swine models, collecting complex hemodynamic data alongside standard toxicology parameters — a data structure notoriously difficult to translate into standard domains.
The approach: Biotech Farm structured surgical and physiological recording forms to align with standard domain definitions from study inception. Baseline vitals, anesthesia depths, and post-operative observations were captured using controlled nomenclature in real time while specialized surgery suites equipped with fluoroscopy and advanced ultrasound recorded cardiovascular endpoints directly into exportable formats.
The result: Technicians populated cardiovascular domains without data loss, and direct electronic recording avoided the transcription errors common when translating unstructured surgical notes into rigid submission formats — protecting both the delivery timeline and data integrity.
Financial and schedule fallout from gateway validation errors
A technical rejection at the submission gateway introduces substantial operational disruption for any commercial sponsor. When the electronic gateway rejects a submission, the agency stops the evaluation clock until a corrected package is uploaded.
Resolving structural metadata errors under urgent deadlines often requires re-contracting external data conversion specialists and pulling scientific staff away from active laboratory programs. A sixty-day delay can compromise corporate financing rounds and push back patent exclusivity windows. Investing in rigorous data hygiene during initial study phases is significantly less expensive than remediating non-compliant files under regulatory scrutiny.
Practical readiness checklist for nonclinical project teams
Research organizations require practical solutions to address technical data conversion requirements during preclinical development. The following breakdown demonstrates how facility capabilities resolve common administrative and regulatory challenges.
| Operational Challenge | Practical Impact | Facility Operational Solution |
|---|---|---|
| Non-standard terminology | Gateway rejection from unmapped vocabulary | Standardized data collection forms linked to CDISC terms |
| Complex surgical endpoints | Loss of quantitative fidelity in device trials | Calibrated surgical imaging tools producing exportable records |
| Data traceability gaps | Reviewer delays during report verification | Integrated data workflows connecting raw data to final reports |
| Unclear trial deviations | Time-consuming agency information requests | Real-time documentation practices supporting reviewer guides |
Establishing these workflows before initiating laboratory studies eliminates conversion bottlenecks and protects project momentum. Rigorous operational controls guarantee that data packages meet international submission criteria.
Preparing sample submissions for preliminary agency evaluation
Sponsors approaching their initial electronic submission can take advantage of agency sample testing programs. The agency provides a validation portal where project teams can upload test packages to verify technical structure before formal filing.
A sample package includes representative domain files, a completed define.xml document, and an accompanying reviewer guide. Submitting this test bundle allows electronic intake engines to run automated validation routines and flag technical discrepancies. Receiving feedback through this voluntary mechanism allows sponsors to correct subtle file naming or structural flaws without risking project timelines.
Frequently asked questions about nonclinical data formatting
Are your preclinical research protocols and study documentation structured to clear electronic regulatory gateways without unexpected project delays? Contact the scientific team at Biotech Farm to discuss how our large animal research facility, advanced surgical platforms, and structured data practices can support your upcoming translational development program.